Licence scope is jurisdiction-specific
An active Canadian medical-device licence demonstrates home-market status; it does not itself authorise placement on the EU market.
Our name goes on the label only after your file stands up to review.
Canadian licensing, MDSAP and ISO 13485 records can support an efficient review. MEDDEV maps them against the separate MDR or IVDR file, EC REP mandate, labelling and EUDAMED responsibilities.
For Canadian medical-device and IVD manufacturers adding the European Union to an established home-market operation.

 Online intake Secure connection



Health Canada issues medical-device licences under the Canadian framework. Union market access remains a separate route, with its own conformity, representative and registration obligations.
An active Canadian medical-device licence demonstrates home-market status; it does not itself authorise placement on the EU market.
MDSAP and ISO 13485 records may support the quality-system review, while MDR or IVDR product obligations still need explicit mapping.
We record owners for the EU declaration, certificates, clinical or performance evidence, labelling, UDI and EUDAMED work before appointment.
Official basis: Health Canada: about medical devices, European Commission: EUDAMED overview, MDR Article 11, European Commission: EUDAMED UDI/Device registration.
Canadian manufacturers may bring Health Canada, MDSAP and ISO 13485 evidence into the review. MEDDEV maps those records to the proposed MDR or IVDR route and checks where the Canadian product family, intended purpose or post-market evidence does not match the planned EU portfolio.
Need the wider comparison? Compare home-market evidence with the EU MDR/IVDR route →
The initial website form contains no document upload. After the parties and scope are confirmed, MEDDEV provides a controlled transfer route for the records below.
Active Medical Device Licence or establishment records relevant to the product scope and the current Canadian label.
MDSAP and ISO 13485 certificates, audit scope and manufacturing-site list.
EU classification rationale, conformity route, declaration draft and notified-body certificates where required.
Technical-documentation index covering risk, clinical or performance evidence, PMS, vigilance and product-liability insurance.
Canadian licence groupings and identifiers may not match the Basic UDI-DI and device families planned for EUDAMED.
Strong Canadian evidence still needs the MDR Annex II and III or corresponding IVDR documentation structure.
The EC REP name and address, EU declarations, UDI data and importer verification package need controlled release.
Four modules — actor registration, UDI/Devices, notified bodies and certificates, and market surveillance — have been mandatory since 28 May 2026 under Commission Decision (EU) 2025/2371 and Regulation (EU) 2024/1860. For devices placed on the market earlier and still being placed on the market, Regulation (EU) 2024/1860 allows no later than twelve months from publication of the Commission notice, which was published on 27 November 2025. Registration data therefore belongs in the market-entry plan, not after it.
MDR Article 10(16) — mirrored for in vitro diagnostics by IVDR Article 10(15) — requires manufacturers to have measures in place providing sufficient financial coverage for potential liability under Directive 85/374/EEC, in a manner proportionate to the risk class, type of device and size of the enterprise. It does not prescribe a particular insurance policy. MEDDEV asks to see those measures because Article 11(5) makes the authorised representative legally liable for defective devices on the same basis as, and jointly and severally with, a manufacturer that has not met its Article 10 obligations. Reviewing the arrangement is our acceptance condition, and that provision is why.
Link MDL or MDEL status, MDSAP records, sites and device families to the proposed EU portfolio.
Record differences in classification, evidence, certificates, declarations, labels and registrations.
Complete file and insurance review, sign the mandate and coordinate the EUDAMED and labelling work.
No. It documents Canadian market status; Union market access follows MDR or IVDR.
They can support the quality-system review, but product-specific MDR or IVDR obligations still require mapping.
MEDDEV reviews the legal manufacturer, intended purpose, classification, identifiers and proposed Basic UDI-DI structure before registration work starts.
We review the arrangement before accepting a mandate, and here is the honest basis for it. MDR Article 10(16) requires measures providing sufficient financial coverage proportionate to the risk class, type of device and size of the enterprise — it does not prescribe a particular policy. Article 11(5) then makes the authorised representative liable for defective devices jointly and severally with a manufacturer that has not met its Article 10 obligations. So the review protects both sides, and it is our acceptance condition rather than a rule we are quoting at you.
MEDDEV accepts only defined device scopes supported by evidence, working safety controls and a manufacturer we can rely on. A commercial enquiry never guarantees appointment. Our name appears on the label only after the acceptance review is complete.
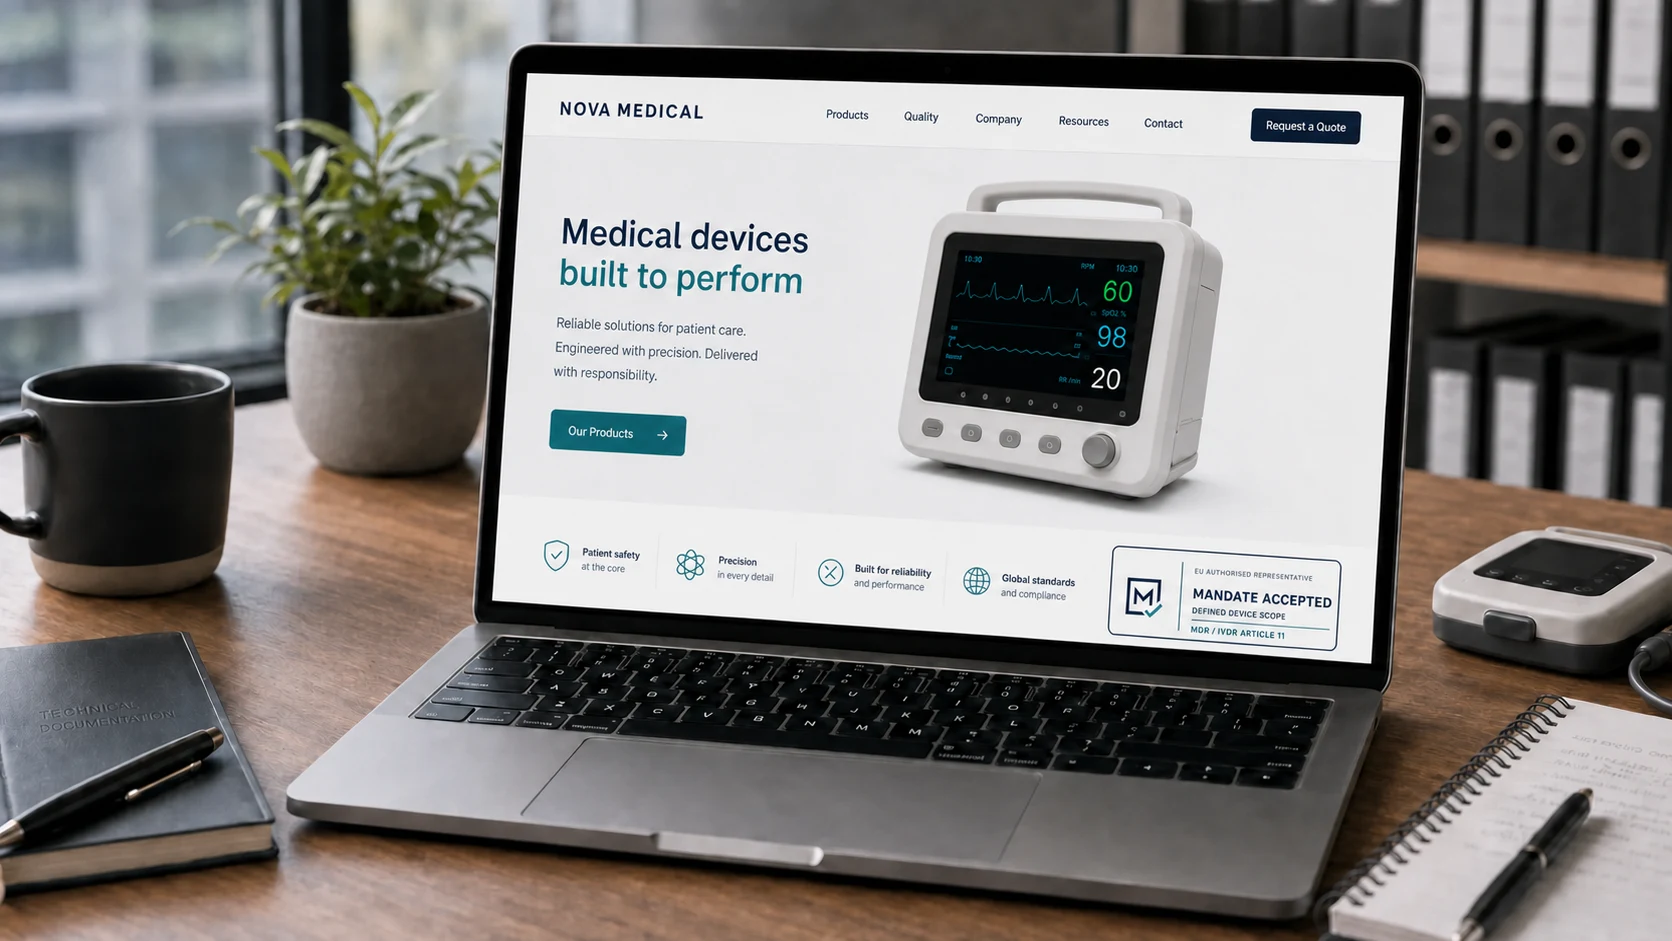
Accepted manufacturers may use the mark where professional buyers assess regulatory readiness. Each use must remain connected to the active MEDDEV mandate and its defined device scope.
Use boundary: the mark must not appear as a CE mark, product certificate or safety approval. It should not be added to the regulated device label, instructions for use or sterile-packaging information panel.
We do not publish a one-size-fits-all price because the mandate depends on the device, portfolio and level of responsibility. The initial review identifies the scope before a fee is proposed.
Classification, IVD or medical device route, number of families, intended purpose and higher-risk features.
Technical documentation, clinical or performance evidence, quality-system records, certificates and labelling.
EUDAMED coordination, vigilance, authority response, importer interface, PRRC support and required review depth.
Fee principle: after the qualification review, we provide a defined scope and a proposal matched to the accepted mandate. No document upload is required in the first contact form.
Confirm the entity, country of establishment, device family and intended EU market.
Have the classification rationale, technical-file index, conformity route and quality evidence available.
Identify complaint, vigilance, field-safety and authority-response owners before the mandate begins.
These anonymised composite scenarios reflect recurring qualification patterns. They are not attributed client case studies or performance claims; the exact scope and outcome depend on the manufacturer and device file.
Problem: home-market approval is mistaken for EU access. Work: map MDR/IVDR, labelling, EUDAMED and importer responsibilities. Typical result: a defined launch route and a documented list of open evidence.
Problem: the outgoing representative controls records and dates are unclear. Work: plan custody, vigilance ownership and handover steps. Typical result: a controlled transition with responsibilities recorded before the new mandate starts.
Problem: a Class IIb, Class III or complex IVD portfolio has unresolved evidence or liability gaps. Work: enhanced file and response-readiness review. Typical result: accept, request defined evidence, or decline before exposure is created.
Each briefing states the provision it relies on and links to the official text. They are written for companies deciding whether, and in what order, to enter the European market.
Four modules became mandatory on 28 May 2026. The twelve-month obligation does not cover every device, and the wording decides which.
Read the EUDAMED briefing →Products covered by the MDR are excluded. Companion software without a medical purpose, portals and service tools are not.
Read the cybersecurity briefing →The notice goes to the authority of the Member State where the manufacturer or its authorised representative is established.
Read the supply briefing →What transfers into a European file, what has no equivalent at home, and where a representative is required by law.
Open the compliance hub →Priority goes to complete, traceable applications with a named regulatory owner. Meeting the entry criteria qualifies a scope for a decision; it does not guarantee acceptance.
The legal entity, authorised signatory, manufacturing sites, device families, models and intended purposes are consistent and traceable.
The qualification, classification and conformity-assessment route are documented, with valid notified-body involvement where required.
The EU declaration, agreed technical-documentation index, certificates, labelling and registration data describe the same device scope.
Risk management, surveillance, complaints, vigilance, recalls and corrective actions have current records, owners and escalation routes.
Named decision-makers can provide records, samples and technical answers and can act within agreed authority-response times.
Insurance, contractual responsibility, document access, sanctions screening, change notification and mandate continuity are acceptable.
What the status means: “MEDDEV Mandate Accepted” confirms MEDDEV’s written acceptance of an EC REP mandate for the defined device scope. It is a trust signal about regulatory readiness and the manufacturer relationship. It is not a CE certificate, product approval, independent safety claim or substitute for the applicable conformity assessment.
Mikołaj Junosza-Szaniawski leads mandate governance, risk oversight and international delivery. Marta Junosza-Szaniawska and Barbara Jęcka-Izdebska remain the named specialists for conformity assessment, registration and ongoing regulatory coordination.
Mikołaj brings more than 25 years of experience at KPMG, including as a Director in Audit Assurance, where he oversaw major sustainability-reporting assurance projects. He combines international project governance with a product-life-cycle perspective encompassing LCA, reuse, recyclability and environmental performance. At MEDDEV, he oversees mandate governance, risk escalation, specialist assignment and continuity of manufacturer communication.
The manufacturer does not need to coordinate a loose group of advisers. The named coordinators define the review scope, assign the relevant specialists and remain accountable for communication and the agreed regulatory work.

Marta coordinates the initial regulatory assessment, defines the file-review scope and confirms whether MEDDEV can accept the representative mandate.
marta.szaniawska@meddev.pl +48 791 480 270
Barbara coordinates registration and notification work and remains a named contact for the practical steps that follow appointment.
barbara.jecka@meddev.pl +48 730 354 190