Great Britain scope
This guide is for manufacturers established in England, Scotland or Wales. It does not assume that a Northern Ireland manufacturer needs an EC REP.
Our name goes on the label only after your file stands up to review.
A manufacturer established in England, Scotland or Wales is outside the European Union. MEDDEV keeps the EU mandate, EUDAMED responsibilities and authority communication distinct from the manufacturer’s Great Britain market records.
For medical-device and IVD manufacturers established in England, Scotland or Wales and maintaining access to the European Union.

 Online intake Secure connection



MHRA guidance separates Great Britain registration from European Union requirements. MEDDEV accepts and manages the EU Article 11 mandate only after reviewing the relevant file and liability scope. Manufacturers established in Northern Ireland require a different analysis.
This guide is for manufacturers established in England, Scotland or Wales. It does not assume that a Northern Ireland manufacturer needs an EC REP.
UK registration records, EU EUDAMED data, certificates and labels need controlled consistency without treating the systems as interchangeable.
Where a manufacturer in Great Britain changes its EC REP, the outgoing and incoming responsibilities are documented to preserve EU regulatory continuity.
Related briefing: The 2025 UK surveillance rules built part of your MDR file already. Where the fifteen-day clock and the periodic report map onto Articles 83 to 88 MDR.
Official basis: MHRA: regulating medical devices in the UK, European Commission: authorised representatives, MDR Article 11, European Commission: EUDAMED UDI/Device registration.
For Great Britain, the practical risk is often the separation between MHRA records and the Union route. We first confirm whether the legal manufacturer is in England, Scotland or Wales, then reconcile the EU portfolio, labels, certificates and EUDAMED responsibilities without treating UK registration as EU authorisation.
Need the wider comparison? Compare home-market evidence with the EU MDR/IVDR route →
The initial website form contains no document upload. After the parties and scope are confirmed, MEDDEV provides a controlled transfer route for the records below.
MHRA manufacturer and device registration details for the Great Britain-established legal manufacturer.
EU declaration draft, MDR or IVDR classification rationale, conformity route and current notified-body certificates.
Technical-documentation index, ISO 13485 certificate, PMS and vigilance procedures and product-liability policy.
Current EC REP mandate and changeover records where the manufacturer is transferring from another representative.
The legal-manufacturer address must establish whether the business is in Great Britain or Northern Ireland before the EU representative need is confirmed.
MHRA records, EU declarations, certificates, labels and EUDAMED data may contain different names, addresses or device groupings.
A transfer needs clear dates, document custody, vigilance ownership and communication with the outgoing representative.
Four modules — actor registration, UDI/Devices, notified bodies and certificates, and market surveillance — have been mandatory since 28 May 2026 under Commission Decision (EU) 2025/2371 and Regulation (EU) 2024/1860. For devices placed on the market earlier and still being placed on the market, Regulation (EU) 2024/1860 allows no later than twelve months from publication of the Commission notice, which was published on 27 November 2025. Registration data therefore belongs in the market-entry plan, not after it.
MDR Article 10(16) — mirrored for in vitro diagnostics by IVDR Article 10(15) — requires manufacturers to have measures in place providing sufficient financial coverage for potential liability under Directive 85/374/EEC, in a manner proportionate to the risk class, type of device and size of the enterprise. It does not prescribe a particular insurance policy. MEDDEV asks to see those measures because Article 11(5) makes the authorised representative legally liable for defective devices on the same basis as, and jointly and severally with, a manufacturer that has not met its Article 10 obligations. Reviewing the arrangement is our acceptance condition, and that provision is why.
Verify the legal-manufacturer address and distinguish Great Britain from Northern Ireland.
Compare MHRA details, certificates, declarations, labels and the planned EU device scope.
Close acceptance gaps, sign or transfer the mandate, then coordinate EC REP labelling and EUDAMED.
No. This page targets manufacturers established in England, Scotland or Wales. Northern Ireland remains subject to a distinct medical-device framework and requires a separate analysis.
No. Great Britain and European Union registrations are separate workstreams.
A UK Responsible Person is the Great Britain role for a manufacturer based outside the UK. This page instead addresses the EU EC REP need of a manufacturer established in Great Britain.
Yes. MEDDEV reviews the changeover terms, document custody, vigilance responsibilities and effective dates before accepting the transfer.
MEDDEV accepts only defined device scopes supported by evidence, working safety controls and a manufacturer we can rely on. A commercial enquiry never guarantees appointment. Our name appears on the label only after the acceptance review is complete.
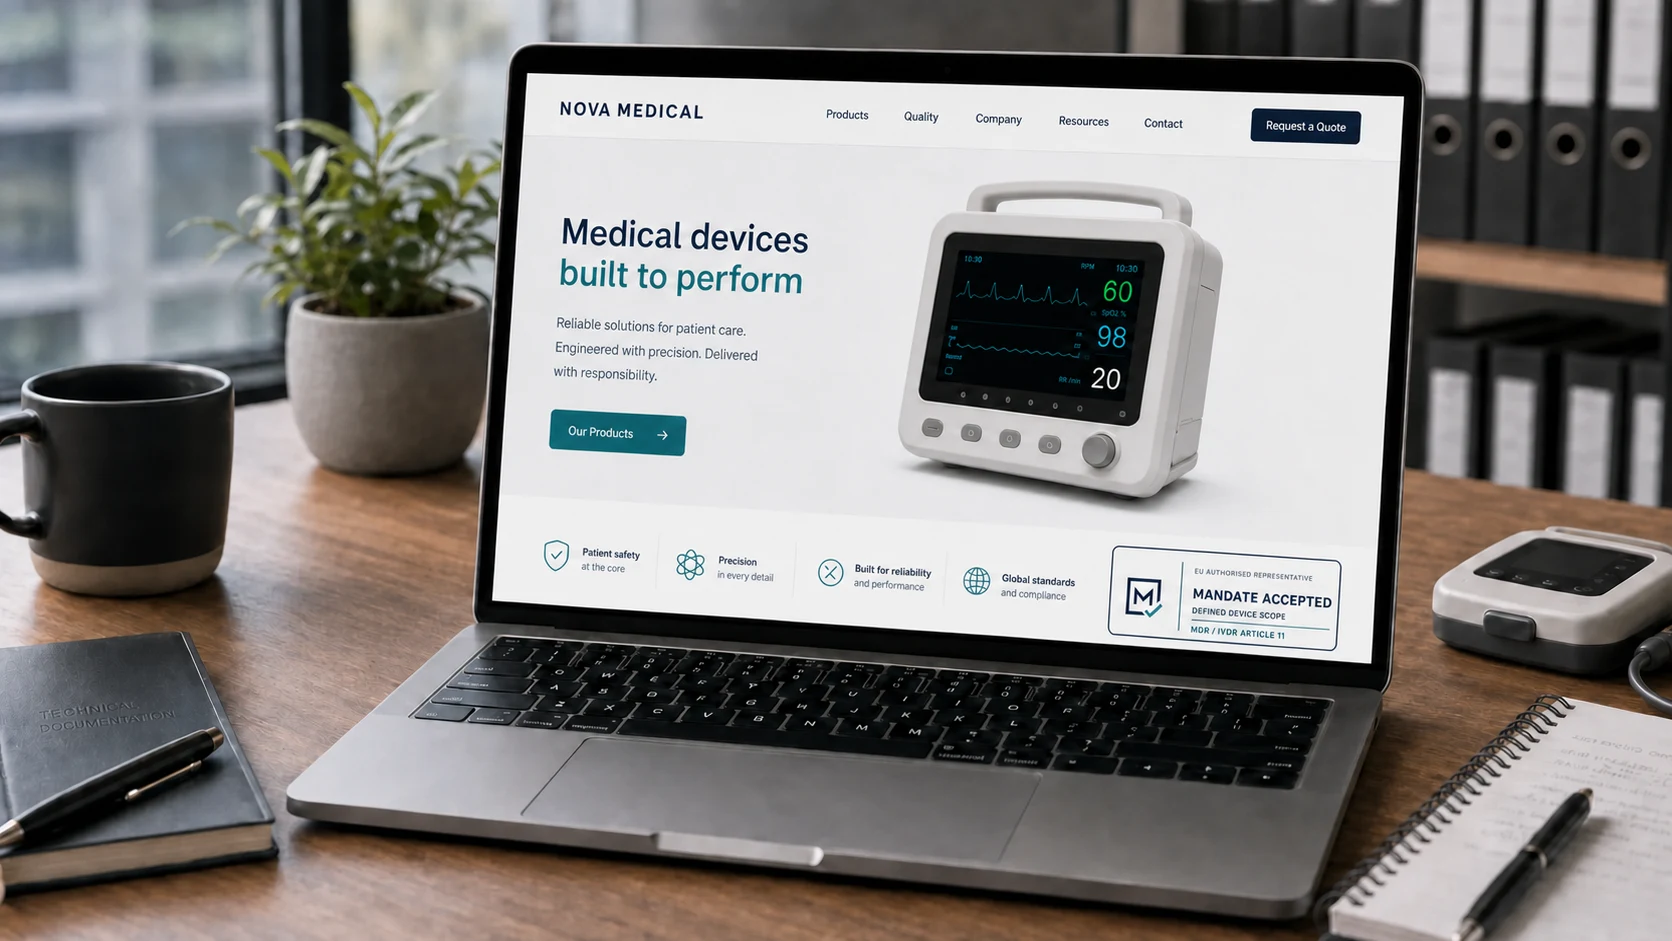
Accepted manufacturers may use the mark where professional buyers assess regulatory readiness. Each use must remain connected to the active MEDDEV mandate and its defined device scope.
Use boundary: the mark must not appear as a CE mark, product certificate or safety approval. It should not be added to the regulated device label, instructions for use or sterile-packaging information panel.
We do not publish a one-size-fits-all price because the mandate depends on the device, portfolio and level of responsibility. The initial review identifies the scope before a fee is proposed.
Classification, IVD or medical device route, number of families, intended purpose and higher-risk features.
Technical documentation, clinical or performance evidence, quality-system records, certificates and labelling.
EUDAMED coordination, vigilance, authority response, importer interface, PRRC support and required review depth.
Fee principle: after the qualification review, we provide a defined scope and a proposal matched to the accepted mandate. No document upload is required in the first contact form.
Confirm the entity, country of establishment, device family and intended EU market.
Have the classification rationale, technical-file index, conformity route and quality evidence available.
Identify complaint, vigilance, field-safety and authority-response owners before the mandate begins.
These anonymised composite scenarios reflect recurring qualification patterns. They are not attributed client case studies or performance claims; the exact scope and outcome depend on the manufacturer and device file.
Problem: home-market approval is mistaken for EU access. Work: map MDR/IVDR, labelling, EUDAMED and importer responsibilities. Typical result: a defined launch route and a documented list of open evidence.
Problem: the outgoing representative controls records and dates are unclear. Work: plan custody, vigilance ownership and handover steps. Typical result: a controlled transition with responsibilities recorded before the new mandate starts.
Problem: a Class IIb, Class III or complex IVD portfolio has unresolved evidence or liability gaps. Work: enhanced file and response-readiness review. Typical result: accept, request defined evidence, or decline before exposure is created.
Each briefing states the provision it relies on and links to the official text. They are written for companies deciding whether, and in what order, to enter the European market.
Four modules became mandatory on 28 May 2026. The twelve-month obligation does not cover every device, and the wording decides which.
Read the EUDAMED briefing →Products covered by the MDR are excluded. Companion software without a medical purpose, portals and service tools are not.
Read the cybersecurity briefing →The notice goes to the authority of the Member State where the manufacturer or its authorised representative is established.
Read the supply briefing →What transfers into a European file, what has no equivalent at home, and where a representative is required by law.
Open the compliance hub →Priority goes to complete, traceable applications with a named regulatory owner. Meeting the entry criteria qualifies a scope for a decision; it does not guarantee acceptance.
The legal entity, authorised signatory, manufacturing sites, device families, models and intended purposes are consistent and traceable.
The qualification, classification and conformity-assessment route are documented, with valid notified-body involvement where required.
The EU declaration, agreed technical-documentation index, certificates, labelling and registration data describe the same device scope.
Risk management, surveillance, complaints, vigilance, recalls and corrective actions have current records, owners and escalation routes.
Named decision-makers can provide records, samples and technical answers and can act within agreed authority-response times.
Insurance, contractual responsibility, document access, sanctions screening, change notification and mandate continuity are acceptable.
What the status means: “MEDDEV Mandate Accepted” confirms MEDDEV’s written acceptance of an EC REP mandate for the defined device scope. It is a trust signal about regulatory readiness and the manufacturer relationship. It is not a CE certificate, product approval, independent safety claim or substitute for the applicable conformity assessment.
Mikołaj Junosza-Szaniawski leads mandate governance, risk oversight and international delivery. Marta Junosza-Szaniawska and Barbara Jęcka-Izdebska remain the named specialists for conformity assessment, registration and ongoing regulatory coordination.
Mikołaj brings more than 25 years of experience at KPMG, including as a Director in Audit Assurance, where he oversaw major sustainability-reporting assurance projects. He combines international project governance with a product-life-cycle perspective encompassing LCA, reuse, recyclability and environmental performance. At MEDDEV, he oversees mandate governance, risk escalation, specialist assignment and continuity of manufacturer communication.
The manufacturer does not need to coordinate a loose group of advisers. The named coordinators define the review scope, assign the relevant specialists and remain accountable for communication and the agreed regulatory work.

Marta coordinates the initial regulatory assessment, defines the file-review scope and confirms whether MEDDEV can accept the representative mandate.
marta.szaniawska@meddev.pl +48 791 480 270
Barbara coordinates registration and notification work and remains a named contact for the practical steps that follow appointment.
barbara.jecka@meddev.pl +48 730 354 190