Sell medical devices in the EU from Türkiye · MDR / IVDR Article 11

EU Authorized Representative for Turkish manufacturers.

Our name goes on the label only after your file stands up to review.

Türkiye is inside the EU–Türkiye Customs Union. In its March 2022 notice to stakeholders the European Commission confirms that a manufacturer established in Türkiye has no obligation to designate an authorised representative in the Union in order to place medical devices on the EU market, and that a device covered by a certificate duly issued by a Türkiye notified body may circulate freely in the EU. An EU representative can still be appointed by choice, and becomes a legal requirement when the legal manufacturer is established outside the EU, the EEA and Türkiye. MEDDEV establishes which case applies before proposing anything.

For Turkish medical-device and IVD manufacturers preparing EU market entry, transferring an existing mandate or coordinating EUDAMED responsibilities.

European Union flag
Established in the European UnionWarsaw, Poland · EC REP under MDR and IVDR Article 11
Online intake Secure connection

Apply for MEDDEV mandate review

A short, three-step intake. We confirm eligibility, define the review scope and decide whether to proceed.

Step 1 of 3 · Manufacturer

No document upload at this stage · reply usually the same working day · in English

Confidential files are handled separately through one-time access, private EU storage and automatic deletion after expiry. Review our security and data handling controls.

MEDDEV brand serving medical-device companies since 2012 MDR 2017/745 & IVDR 2017/746 Mandate accepted only after review
Manufacturers who have worked with MEDDEV
CHM Sp. z o.o.MDD Sp. z o.o.WDI Wise Device EuropeGlobal Pharma CMSensiMedUSP ZdrowieAdamed Consumer HealthcarePiórex S.A.
Türkiye → European Union

The Customs Union changes the question before it changes the paperwork.

Most guidance written for third-country manufacturers does not apply to Türkiye. Decisions Nos 1/95, 2/97 and 1/2006 of the EC–Türkiye Association Council, together with the Joint Committee statements of 21 May 2021 and 13 September 2021, confirm that Turkish legislation is aligned with Regulations (EU) 2017/745 and 2017/746, and that references to the territories of the Parties are read as the territory of the Customs Union. MEDDEV starts by establishing which side of that line the legal manufacturer actually sits on.

No obligation to appoint an EU representative

The Commission states that manufacturers established in Türkiye have no obligation to designate an authorised representative on EU territory in order to place medical devices on the EU market. The reverse is also true: EU manufacturers need no representative in Türkiye. Any provider telling a Turkish manufacturer that an EC REP is legally required is misreading the Customs Union.
A Turkish notified body carries EU weight

Türkiye is entitled to designate notified bodies under Regulations 2017/745 and 2017/746, and under Decision No 1/2006 may designate more than one. A notified body established in Türkiye has the same rights and obligations as one established in the EU, and devices covered by a certificate it duly issued may circulate freely in the EU.

The exception is where the legal manufacturer sits

A manufacturer established outside the EU, the EEA and Türkiye designates one authorised representative — in the EU or in Türkiye — to place devices on either market. Group structures, contract manufacturing and own-brand labelling are where this is most often misjudged, because the entity on the label is not always the entity established in Türkiye.

Related briefing: Your ÜTS records answer most of the questions EUDAMED asks. Which device data maps across, and which level is usually missing.

Official basis: Türkiye ÜTS: medical-device product tracking system, European Commission: EU–Türkiye Customs Union and medical devices, European Commission: authorised representatives, MDR Article 11, European Commission: EUDAMED UDI/Device registration.

Local review focus

What usually needs attention for Türkiye manufacturers.

For manufacturers in Türkiye, the EU–Türkiye relationship can create confusion between customs arrangements and medical-device conformity. MEDDEV separates the customs and distribution question from the MDR or IVDR route, then verifies the legal manufacturer, device scope and applicable EUDAMED responsibilities.

Qualification file

What to prepare for the first EU review.

The initial website form contains no document upload. After the parties and scope are confirmed, MEDDEV provides a controlled transfer route for the records below.

  1. 01

    The legal manufacturer’s place of establishment, evidenced rather than assumed: registration details, signatory powers and the entity named on the label and declaration.

  2. 02

    Certificates issued by a notified body, with the designation checked against the current NANDO entry for the body and the procedure.

  3. 03

    EU classification rationale, intended purpose and the conformity route the certificate actually covers.

  4. 04

    ÜTS records for the device families concerned, and product-liability insurance covering the Union market.

Typical transition gaps

Issues to resolve before the mandate starts.

Representation sold as an obligation

Turkish manufacturers are routinely offered EC REP mandates as a legal requirement. Where the legal manufacturer is established in Türkiye it is not one. A mandate may still be worth having — for a single EU-facing contact, or to coordinate EUDAMED and vigilance work — but it is a commercial decision, not a legal necessity. Note that appointing a representative under Article 11 is not a light arrangement: it carries a defined statutory mandate, and the registration obligations themselves remain the manufacturer’s.

The label entity and the established entity differ

Whether a representative is needed follows the place of establishment of the legal manufacturer — not the shipping address, the group headquarters or where the goods clear customs. Where the entity named on the declaration is established outside the EU, the EEA and Türkiye, one representative must be designated.

Certificate scope narrower than the portfolio

A certificate supports only what it actually covers, and only while it remains valid. MDR certificates run for a defined period and can be suspended, restricted or withdrawn. Device families added after certification, or intended purposes broadened for the EU market, fall outside it.

EUDAMED is no longer a future obligation

Four modules — actor registration, UDI/Devices, notified bodies and certificates, and market surveillance — have been mandatory since 28 May 2026 under Commission Decision (EU) 2025/2371 and Regulation (EU) 2024/1860. For devices placed on the market earlier and still being placed on the market, Regulation (EU) 2024/1860 allows no later than twelve months from publication of the Commission notice, which was published on 27 November 2025. Registration data therefore belongs in the market-entry plan, not after it.

Financial coverage: what the law says, and what we ask for

MDR Article 10(16) — mirrored for in vitro diagnostics by IVDR Article 10(15) — requires manufacturers to have measures in place providing sufficient financial coverage for potential liability under Directive 85/374/EEC, in a manner proportionate to the risk class, type of device and size of the enterprise. It does not prescribe a particular insurance policy. MEDDEV asks to see those measures because Article 11(5) makes the authorised representative legally liable for defective devices on the same basis as, and jointly and severally with, a manufacturer that has not met its Article 10 obligations. Reviewing the arrangement is our acceptance condition, and that provision is why.

Country-specific route

From home-market records to EU representation.

01

Establish the position

Confirm where the legal manufacturer is established and whether the Customs Union rules apply. This determines whether representation is a legal obligation, a commercial option or irrelevant.

02

Test the certificate and the scope

Check the notified body’s designation, the procedures it covers and whether the certified scope matches the portfolio intended for the Union market.

03

Decide, then document

Where a mandate makes sense, MEDDEV accepts it after the usual file, insurance and sanctions review. Where it does not, we say so and set out what actually needs attention.

Check this portfolio →
Questions from Türkiye

What manufacturers ask before appointment.

Does ÜTS registration authorise EU sales?

No. ÜTS is the Turkish product tracking system and records local-market status. Union market access follows the conformity assessment and the certificate, which under the Customs Union may have been issued by a Turkish or an EU notified body.

Is a certificate from a Turkish notified body accepted in the EU?

Yes, where the body is designated for the relevant Regulation and the relevant procedure and device codes. The Commission states that a notified body established in Türkiye has the same rights and obligations as one established in the EU, and that devices covered by a certificate duly issued by it may circulate freely in the EU. Two things need checking, not one: the body’s current designation in NANDO, and the certificate’s own validity and status. A certificate is also not a market authorisation — the declaration of conformity, CE marking, labelling, UDI and the applicable EUDAMED entries still have to be in place.

We are a Turkish company but the legal manufacturer is our parent abroad. What then?

Then the Customs Union position does not help you. A manufacturer established outside both the EU and Türkiye must designate one authorised representative, in the EU or in Türkiye. This is the single most common misreading we see, and it is worth settling before anything is printed on a label.

Does a manufacturer in Türkiye need an EC REP?

Not as a legal obligation. The European Commission states that manufacturers established in Türkiye have no obligation to designate an authorised representative on EU territory in order to place medical devices on the EU market, because Türkiye lies within the Customs Union. Where the legal manufacturer is established outside the EU, the EEA and Türkiye, one representative is designated — in the EU or in Türkiye. MEDDEV confirms where the legal manufacturer is established before any mandate is discussed, and will say so plainly if a mandate is not required.

What is reviewed before the mandate?

MEDDEV reviews the legal manufacturer, device identity, classification, conformity route, technical-documentation index, insurance, vigilance arrangements and sanctions screening.

Can documents be sent through the first contact form?

No. The form is for qualification only. A controlled document-transfer route is provided after scope and parties are confirmed.

Post-project client survey

Post-project ratings from real regulatory work.

56published post-project ratings
97%marked MEDDEV “recommended”
5/5 across five criteria WDI Wise Device Europe Readiness audit · 2015
5/5 across five criteria SensiMed Medical device notification · 2015
5/5 across five criteria Piórex S.A. Medical device registration · 2015

Historical MEDDEV post-project survey across regulatory services. Clients rated knowledge transfer, consultant engagement, subject-matter preparation, problem solving and overall service. The individual ratings shown here are from 2015; the summary figures cover the published set from that survey programme. Comments were optional, so most ratings carry none — and no quotation has been reconstructed or written on a client’s behalf.

Acceptance is earned, not purchased

Not every mandate is accepted.

MEDDEV accepts only defined device scopes supported by evidence, working safety controls and a manufacturer we can rely on. A commercial enquiry never guarantees appointment. Our name appears on the label only after the acceptance review is complete.

MEDDEV mandate reviewAcceptance gate
Manufacturer, device and intended purposeAcceptance criterion
Conformity route and regulatory evidenceAcceptance criterion
Safety, post-market and vigilance controlsAcceptance criterion
Communication, response and liability arrangementsAcceptance criterion
Outcome: mandate accepted, conditions issued or mandate declined.
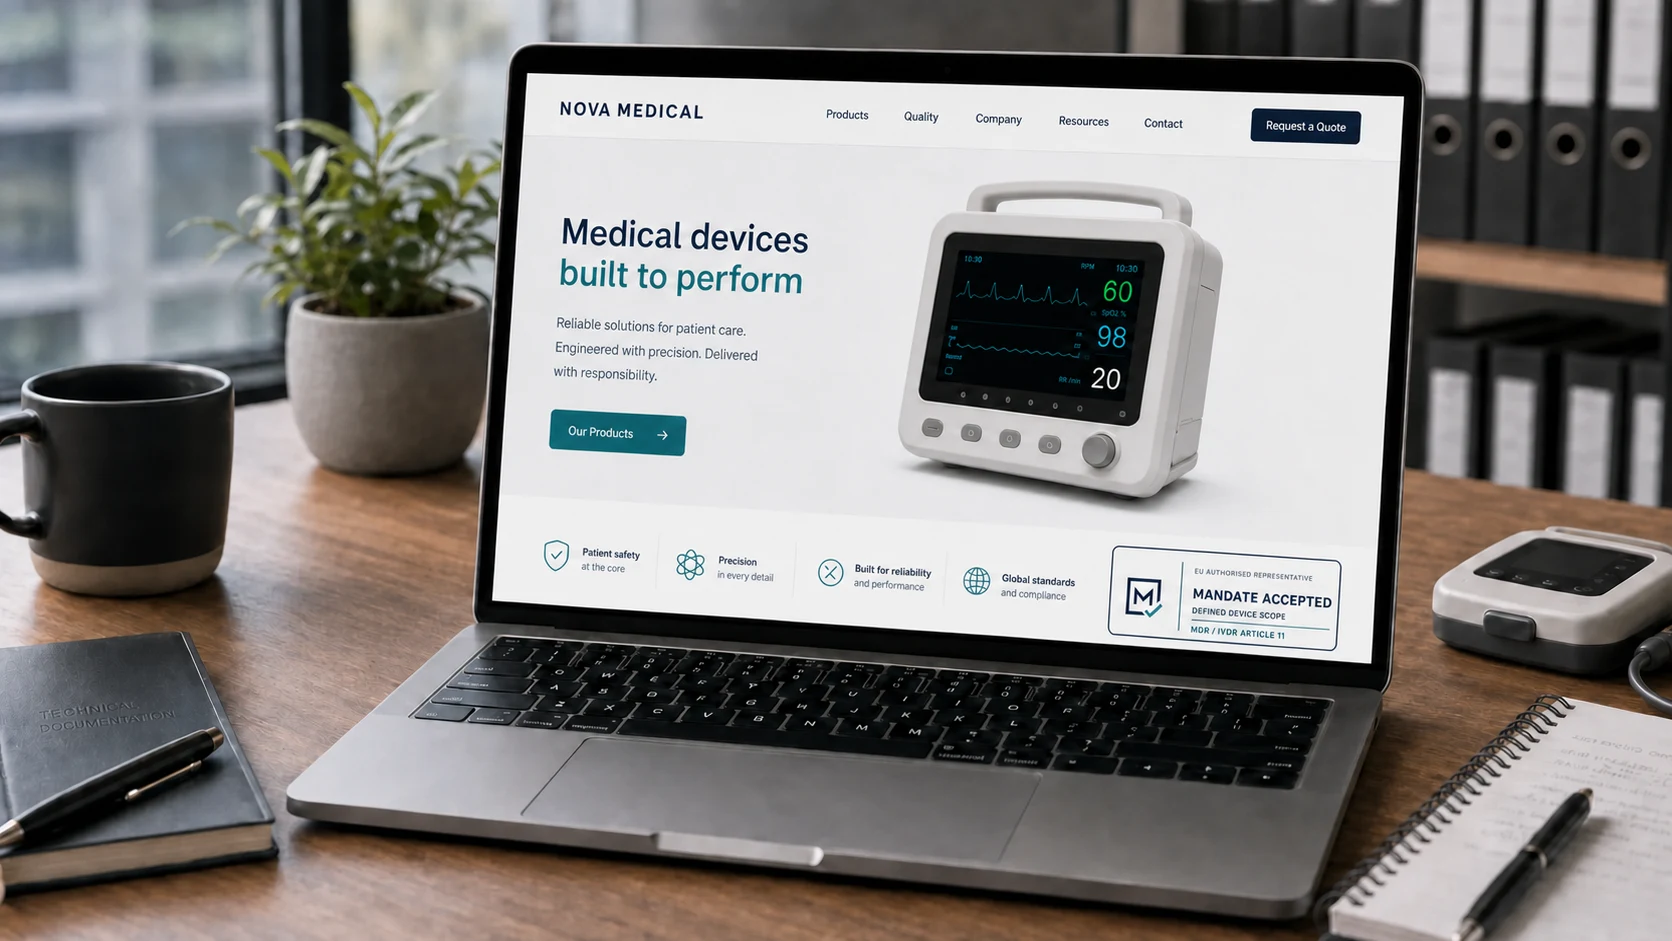
Example of the MEDDEV Mandate Accepted mark displayed on a medical-device manufacturer's website
Example on a manufacturer website. The issued mark should link to the active MEDDEV mandate and its defined device scope.
A status worth displaying

Turn regulatory acceptance into a visible trust signal.

Accepted manufacturers may use the mark where professional buyers assess regulatory readiness. Each use must remain connected to the active MEDDEV mandate and its defined device scope.

  • Manufacturer websiteAlongside EU market-access information and the represented device portfolio.
  • Product catalogueOn pages covering devices included in the accepted mandate.
  • Distributor dossierAs a verifiable representation status during onboarding and due diligence.
  • Sales presentationFor procurement, partner meetings and trade-fair communication.

Use boundary: the mark must not appear as a CE mark, product certificate or safety approval. It should not be added to the regulated device label, instructions for use or sterile-packaging information panel.

Before you enquire

Know what shapes the review and the fee.

We do not publish a one-size-fits-all price because the mandate depends on the device, portfolio and level of responsibility. The initial review identifies the scope before a fee is proposed.

Device and portfolio

Classification, IVD or medical device route, number of families, intended purpose and higher-risk features.

Evidence readiness

Technical documentation, clinical or performance evidence, quality-system records, certificates and labelling.

Ongoing responsibility

EUDAMED coordination, vigilance, authority response, importer interface, PRRC support and required review depth.

Fee principle: after the qualification review, we provide a defined scope and a proposal matched to the accepted mandate. No document upload is required in the first contact form.

01

Legal manufacturer

Confirm the entity, country of establishment, device family and intended EU market.

02

Core evidence

Have the classification rationale, technical-file index, conformity route and quality evidence available.

03

Post-market readiness

Identify complaint, vigilance, field-safety and authority-response owners before the mandate begins.

Check your EC REP readiness →
Typical review scenarios

The questions we resolve before a mandate starts.

These anonymised composite scenarios reflect recurring qualification patterns. They are not attributed client case studies or performance claims; the exact scope and outcome depend on the manufacturer and device file.

01

New EU launch

Problem: home-market approval is mistaken for EU access. Work: map MDR/IVDR, labelling, EUDAMED and importer responsibilities. Typical result: a defined launch route and a documented list of open evidence.

02

Change of EC REP

Problem: the outgoing representative controls records and dates are unclear. Work: plan custody, vigilance ownership and handover steps. Typical result: a controlled transition with responsibilities recorded before the new mandate starts.

03

Higher-risk review

Problem: a Class IIb, Class III or complex IVD portfolio has unresolved evidence or liability gaps. Work: enhanced file and response-readiness review. Typical result: accept, request defined evidence, or decline before exposure is created.

Regulatory briefings

The obligations manufacturers outside the Union ask about most.

Each briefing states the provision it relies on and links to the official text. They are written for companies deciding whether, and in what order, to enter the European market.

EUDAMED · Article 123(3)

Device data falls due on 27 November 2026.

Four modules became mandatory on 28 May 2026. The twelve-month obligation does not cover every device, and the wording decides which.

Read the EUDAMED briefing →
Cyber Resilience Act

Reporting duties start on 11 September 2026.

Products covered by the MDR are excluded. Companion software without a medical purpose, portals and service tools are not.

Read the cybersecurity briefing →
Article 10a MDR

Withdrawing a device now takes six months of notice.

The notice goes to the authority of the Member State where the manufacturer or its authorised representative is established.

Read the supply briefing →
Manufacturers outside the EU

Not every part of a domestic file has to be rebuilt.

What transfers into a European file, what has no equivalent at home, and where a representative is required by law.

Open the compliance hub →
Published acceptance criteria

Six tests every device scope must pass.

Priority goes to complete, traceable applications with a named regulatory owner. Meeting the entry criteria qualifies a scope for a decision; it does not guarantee acceptance.

  1. 01

    Verified legal manufacturer and scope

    The legal entity, authorised signatory, manufacturing sites, device families, models and intended purposes are consistent and traceable.

  2. 02

    Defensible EU regulatory route

    The qualification, classification and conformity-assessment route are documented, with valid notified-body involvement where required.

  3. 03

    Controlled regulatory evidence

    The EU declaration, agreed technical-documentation index, certificates, labelling and registration data describe the same device scope.

  4. 04

    Working safety and post-market controls

    Risk management, surveillance, complaints, vigilance, recalls and corrective actions have current records, owners and escalation routes.

  5. 05

    Responsive manufacturer relationship

    Named decision-makers can provide records, samples and technical answers and can act within agreed authority-response times.

  6. 06

    Liability and continuity secured

    Insurance, contractual responsibility, document access, sanctions screening, change notification and mandate continuity are acceptable.

What the status means: “MEDDEV Mandate Accepted” confirms MEDDEV’s written acceptance of an EC REP mandate for the defined device scope. It is a trust signal about regulatory readiness and the manufacturer relationship. It is not a CE certificate, product approval, independent safety claim or substitute for the applicable conformity assessment.

Present my device scope →
Meet MEDDEV in your own time zone. Live reference times for manufacturer markets. MEDDEV’s registered office is in Warsaw.
Request a 30-minute online meeting
Vancouver–:–
New York / Toronto–:–
Warsaw–:–
Dubai–:–
Tokyo / Seoul–:–
Sydney–:–
The people behind your mandate

Director-level governance. Named specialists for regulatory execution.

Mikołaj Junosza-Szaniawski leads mandate governance, risk oversight and international delivery. Marta Junosza-Szaniawska and Barbara Jęcka-Izdebska remain the named specialists for conformity assessment, registration and ongoing regulatory coordination.

MEDDEV Compliance sp. z o.o. · Warsaw, European Union ul. Mydlarska 47, 04-690 Warszawa, Poland biuro@meddev.pl
Check my eligibility →
Mikołaj Junosza-Szaniawski, Managing Director of MEDDEV EC REP operations and governance

Mikołaj Junosza-Szaniawski

Managing Director · EC REP Operations & Governance

Mikołaj brings more than 25 years of experience at KPMG, including as a Director in Audit Assurance, where he oversaw major sustainability-reporting assurance projects. He combines international project governance with a product-life-cycle perspective encompassing LCA, reuse, recyclability and environmental performance. At MEDDEV, he oversees mandate governance, risk escalation, specialist assignment and continuity of manufacturer communication.

25+ years at KPMG Director, Audit Assurance MBA · Carlson School of Management Specialist LCA training · MIT ESG lecturer · Kozminski University Programme manager · EPD Polska
One accountable contact, specialist depth when the device requires it.

The manufacturer does not need to coordinate a loose group of advisers. The named coordinators define the review scope, assign the relevant specialists and remain accountable for communication and the agreed regulatory work.

Regulatory affairsQuality systemsClinical evaluationPhysiciansBiomedical engineeringSoftware and cybersecurityAesthetic devicesPost-market surveillance
Marta Junosza-Szaniawska, MEDDEV conformity assessment specialist

Marta Junosza-Szaniawska

Conformity assessment & EC REP mandate review

Marta coordinates the initial regulatory assessment, defines the file-review scope and confirms whether MEDDEV can accept the representative mandate.

marta.szaniawska@meddev.pl +48 791 480 270
Barbara Jęcka-Izdebska, MEDDEV registration and notification specialist

Barbara Jęcka-Izdebska

Registration, notifications & ongoing coordination

Barbara coordinates registration and notification work and remains a named contact for the practical steps that follow appointment.

barbara.jecka@meddev.pl +48 730 354 190
Check eligibility →
WhatsApp
Scroll to Top